
 English
 Italian
 Spanish
 Thai
 Japanese
 Portuguese
 Korean
 Russian
 French
 German
 Polish
 Arabic
 Indonesian

For pharmaceutical chemical sourcing, a high assay result is only one part of material quality. Related substances, residual solvents, water content, and batch-to-batch variation can all influence downstream manufacturing performance. When evaluating a Crotamitone pharmaceutical intermediate supplier China, it is therefore important to understand how the manufacturer controls impurities throughout production rather than relying exclusively on the final Certificate of Analysis. Effective quality management begins with raw material qualification, continues through reaction and purification, and ends with analytical release and batch traceability.
Why Impurity Control Is Essential
Impurities in pharmaceutical chemicals can originate from several sources. They may be introduced through starting materials, generated during chemical reactions, formed by side reactions, or produced during purification and storage. Their presence and concentration depend on the manufacturing route, process conditions, raw material quality, and handling environment.
For Crotamitone-related materials, the applicable impurity profile should be established according to the exact material specification and its intended manufacturing stage. An upstream intermediate and a finished API should not automatically share the same acceptance criteria. The downstream process may remove certain impurities, while others may be difficult to purge or could affect subsequent reaction performance.
This is why a supplier should understand not only the target assay but also the impurity behavior of the manufacturing process. A stable process aims to minimize unwanted by-products at their source, making purification more predictable and reducing the risk of batch rejection.
Raw Material Qualification and Incoming Inspection
Raw material quality is the starting point for consistent pharmaceutical chemical production. Variations in starting material purity, moisture, composition, or trace impurities may influence reaction conversion and generate different impurity patterns in the finished batch.
A controlled incoming inspection process should define supplier qualification requirements, material specifications, sampling procedures, and acceptance criteria. Critical raw materials may require identity testing and additional analysis based on their risk profile. Supplier changes should be reviewed before implementation because a different manufacturing source may introduce variations that are not immediately visible from a basic specification.
Traceability is equally important. Each raw material batch should be linked to the production batch in which it is used. If an unexpected impurity appears during finished-product testing, this information helps the quality team investigate whether the issue is associated with a specific raw material lot or a manufacturing-stage deviation.
For long-term supply arrangements, consistency in raw material sourcing can support more predictable production. However, maintaining an approved supplier list should not prevent periodic supplier performance reviews, especially when changes in raw material origin, specifications, or manufacturing processes occur.
Process Control and the Formation of Related Substances
Reaction conditions are among the most important factors influencing impurity formation. Temperature, pressure, reaction time, reagent ratio, concentration, and mixing efficiency can affect both conversion and selectivity. If process parameters drift outside their established ranges, the reaction may generate additional by-products or leave higher levels of unreacted starting materials.
A well-designed manufacturing process identifies critical process parameters and defines suitable operating ranges. In high-temperature or high-pressure operations, accurate instrumentation and equipment controls are particularly important. Hydrogenation processes also require appropriate control of reaction conditions and safety systems. These capabilities matter because process reliability depends on the interaction between chemistry, equipment design, and operating discipline.
Purification must then address the impurities relevant to the material specification. Distillation, crystallization, filtration, washing, or other separation techniques may be used depending on the compound and manufacturing route. High-vacuum distillation, for example, can support separation of suitable compounds under reduced pressure, but its applicability depends on the material's thermal stability, volatility, and process characteristics. No single purification technique is appropriate for every chemical system.
Jiangsu Jingye Pharmaceutical has manufacturing capabilities in high-temperature and high-pressure reactions, hydrogenation, chiral synthesis, and high-vacuum distillation. These capabilities support a range of pharmaceutical chemical manufacturing processes. The specific process applied to a Crotamitone-related product should be confirmed through product-level technical documentation and supplier communication.
Analytical Testing: What Should a CoA Demonstrate?
The Certificate of Analysis is a key document for evaluating a pharmaceutical chemical batch, but its value depends on the relevance and reliability of the reported tests. A useful CoA should clearly identify the product, batch number, manufacturing or testing date as applicable, specification, test method, result, and release status.
Assay testing measures the content of the target compound according to the specified method. Related-substance testing evaluates relevant impurities and degradation products. HPLC is commonly used for these purposes when the analytical method is suitable for the compound and intended application. Identification may involve techniques such as infrared spectroscopy or other appropriate methods, depending on the specification. Water content and residual solvent testing may also be required when relevant to product quality and downstream use.
The method itself deserves attention. Different analytical methods can produce different results if sample preparation, chromatographic conditions, reference standards, or integration rules are inconsistent. A supplier should have documented analytical procedures and suitable reference materials. Where customer specifications differ from the supplier's routine methods, both parties should agree on the testing approach and acceptance criteria before the first commercial batch.
For critical applications, method transfer or comparative testing may be necessary. This is especially relevant when a customer plans to compare supplier results with internal laboratory data. A clear agreement on analytical methods helps prevent disputes caused by differences in testing conditions rather than actual product quality.
Batch-to-Batch Consistency and Change Management
A single conforming batch does not establish long-term process reliability. Pharmaceutical manufacturers need confidence that successive batches will meet the same agreed specification. Batch consistency should therefore be assessed through historical CoAs, production records where available, deviation trends, and supplier quality performance.
A supplier should maintain a formal change-control process for changes that could affect product quality. Examples include adjustments to raw material sources, manufacturing equipment, process parameters, analytical methods, production sites, and packaging systems. Customers should be informed of relevant changes according to the agreed quality and supply arrangements.
Deviation management is another important indicator of manufacturing maturity. When a batch fails an acceptance criterion or an unexpected result occurs, the manufacturer should investigate the root cause, assess the impact on other batches, and implement appropriate corrective and preventive actions. Simply repeating a test without investigating the cause is not an adequate substitute for a documented quality investigation.
Storage, Packaging, and Material Stability
Quality control does not end when a batch passes final testing. Packaging and storage conditions can influence chemical stability during transportation and warehousing. Moisture exposure, temperature variation, light, and incompatible packaging materials may affect product quality depending on the compound's characteristics.
Packaging should be selected based on the product's stability data and shipping requirements. Buyers should confirm the recommended storage conditions, shelf life or retest period, packaging configuration, and handling precautions. For international shipments, packaging integrity and clear batch identification are particularly important because materials may pass through multiple logistics stages before reaching the manufacturing site.
A supplier should provide appropriate handling information through the product specification and SDS. If special storage or transportation conditions apply, these should be communicated before shipment rather than after the material arrives.
Working with a Quality-Oriented Manufacturer in China
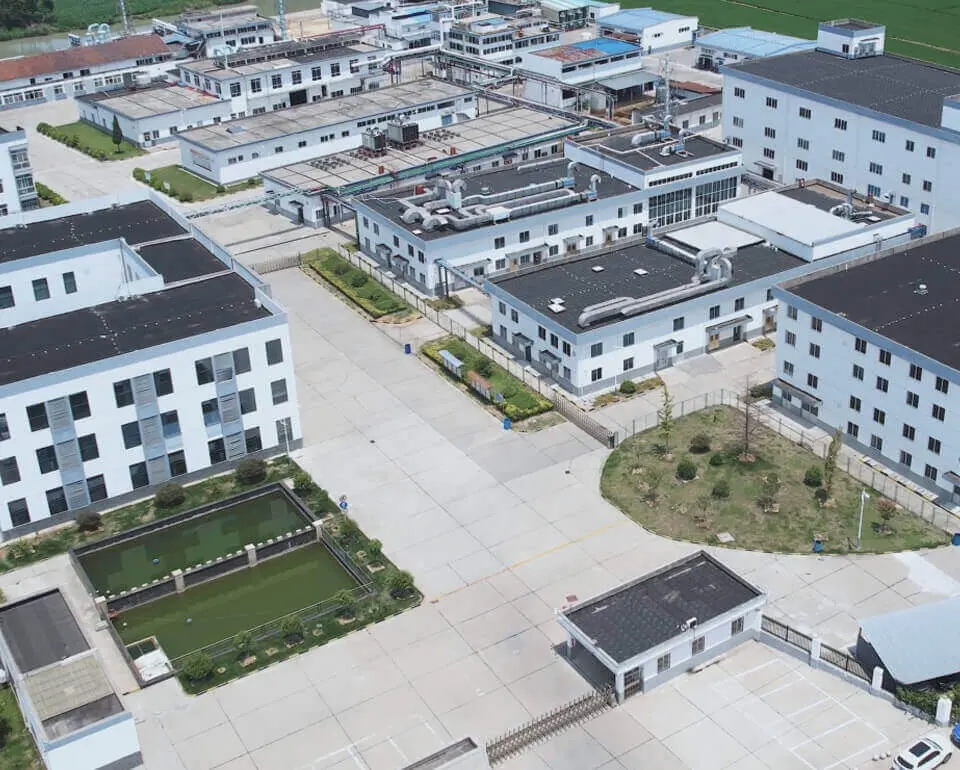
Jiangsu Jingye Pharmaceutical, founded in 1994 and restructured in 2016, integrates R&D, production, and international trade. Its manufacturing site in Changzhou covers approximately 50,000 square meters and is supported by a professional technical team, modern production facilities, and analytical instruments. The company reports operating in accordance with GMP standards and holding a Drug Production License, ISO 9001, ISO 14001, and GB/T 45001 certifications.
Its quality management approach is supported by production controls and an EHS management system. Cooperation with universities and research institutions also provides a foundation for ongoing technical development. For international customers, the company has established export relationships across Europe, North America, Southeast Asia, Japan, and South Korea.
Building a More Reliable Crotamitone Supply Process
Choosing a Crotamitone pharmaceutical intermediate supplier China requires a systematic review of impurity control, analytical methods, process consistency, and change management. Buyers should request product-specific specifications and representative CoAs, confirm relevant testing methods, and assess whether the supplier can maintain consistent quality across repeat orders.
The most reliable sourcing arrangement is built on transparent specifications and documented quality responsibilities. When the supplier and customer agree on material identity, impurity limits, analytical methods, packaging, and change-control expectations before commercial production, the risk of downstream quality problems becomes easier to manage.