
 English
 Italian
 Spanish
 Thai
 Japanese
 Portuguese
 Korean
 Russian
 French
 German
 Polish
 Arabic
 Indonesian

How to Evaluate Product Quality, Analytical Data, and Batch Consistency
When evaluating a 2-Bromo-1-[4-(methylsulfonyl)phenyl]-1-ethanone pharmaceutical intermediate supplier China, product quality should be assessed through more than a single purity result. This compound is an organic brominated aromatic ketone containing a methylsulfonyl substituent, and its suitability for pharmaceutical development or subsequent manufacturing depends on its identity, impurity profile, analytical traceability, and consistency between batches. A supplier that can provide reliable analytical documentation and a controlled quality system helps reduce the risk of unexpected results during downstream processing.
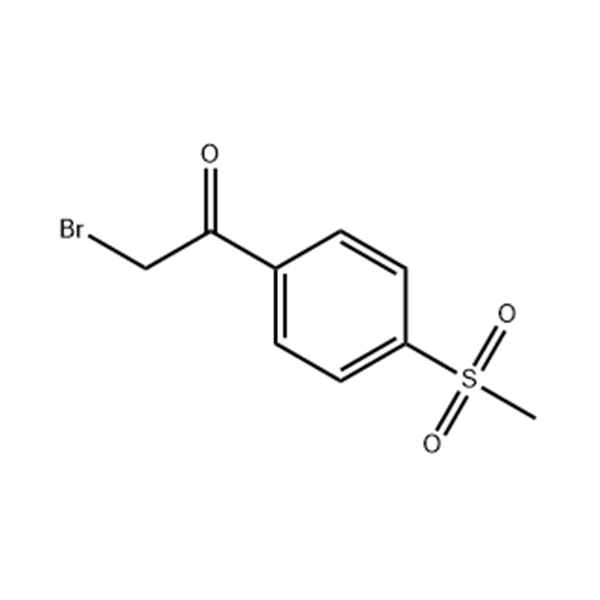
For pharmaceutical intermediate sourcing, the key question is not simply whether a material meets a stated specification. It is whether the supplier can demonstrate that the specification is technically appropriate, the analytical methods are suitable, and each released batch can be traced to its supporting quality records. These considerations become particularly important when a material moves from initial evaluation to process validation, routine manufacturing, or international supply.
Understanding the Product and Its Quality Requirements
2-Bromo-1-[4-(methylsulfonyl)phenyl]-1-ethanone is also commonly described by the structural name 2-bromo-1-[4-(methylsulfonyl)phenyl]ethan-1-one. Its molecular structure includes an aromatic ring, a methylsulfonyl group, a carbonyl group, and a brominated carbon adjacent to the carbonyl functionality. These structural features determine the compound's chemical identity and influence the analytical questions that should be addressed during material qualification.
The molecular formula is C₉H₉BrO₃S, with a theoretical average molecular weight of approximately 277.14 g/mol. Such information provides a starting point for confirming identity, but molecular formula and molecular weight alone are not sufficient to establish material quality. A complete assessment should connect structural confirmation with quantitative purity, related substances, and other attributes defined in the agreed product specification.
For a pharmaceutical intermediate, the required specification should be established according to its intended use and the customer's process requirements. A material intended for early-stage research may be evaluated differently from one intended for a controlled commercial manufacturing process. The distinction should be documented rather than assumed, particularly when the same product is offered to customers working under different quality systems.
Why Purity Alone Is Not Enough
A certificate of analysis may report a high chromatographic purity result, but that figure needs to be interpreted within the context of the analytical method. For example, an HPLC area percentage represents the relative detector response of detected chromatographic peaks under specified conditions. It does not automatically equal the mass percentage of the target compound, nor does it establish the absence of impurities that the method cannot adequately detect.
A technically useful quality package should identify the analytical method, the result, the acceptance criterion, and the applicable batch number. If the method is based on HPLC, the supporting information should make it possible to understand how the target peak is identified and how relevant impurities are evaluated. Where appropriate, method validation or verification records can provide additional evidence of analytical suitability.
Identity testing should be considered separately from purity testing. Depending on the agreed specification, techniques such as NMR, mass spectrometry, or infrared spectroscopy may support structural confirmation. HPLC can provide valuable information about chromatographic composition, while other methods may address questions that chromatography alone cannot resolve. The exact combination should reflect the compound, the risk assessment, and the customer's quality requirements.
This distinction matters when comparing quotations from different suppliers. Two materials with the same stated purity may not have equivalent impurity profiles, documentation quality, or analytical confidence. Comparing only the headline purity number can therefore create a misleading impression of equivalence.
Impurity Profiling and Downstream Process Compatibility
The impurity profile of a pharmaceutical intermediate can influence the performance of subsequent processing steps. Depending on the downstream chemistry, trace impurities may affect reaction selectivity, product isolation, purification efficiency, or the impurity profile of a later intermediate. The actual impact depends on the customer's process and should be established through technical evaluation rather than generalized assumptions.
For 2-Bromo-1-[4-(methylsulfonyl)phenyl]-1-ethanone, the supplier and customer should agree on the relevant impurity controls based on the manufacturing route, available analytical knowledge, and intended application. Potential related substances should be evaluated using scientifically justified methods. Where a specific impurity is known to be relevant, a defined limit and an appropriate analytical approach may be more useful than an undifferentiated total-purity requirement.
Residual solvents, water content, inorganic residues, and other quality attributes may also require consideration when relevant to the manufacturing process or material specification. Not every attribute needs to be tested for every application. The objective is to establish a proportionate control strategy that addresses actual risks and provides meaningful evidence of suitability.
A supplier's response to technical questions is also informative. Can the supplier explain which analytical methods support the reported results? Can it clarify the distinction between a routine release test and an additional customer-specific test? Can it assess a proposed specification before accepting an order? These capabilities help determine whether quality control is integrated into manufacturing or treated merely as a final documentation exercise.
Batch Traceability and Quality Documentation
Batch traceability is essential when an intermediate is used in a regulated or tightly controlled manufacturing environment. Each shipment should be linked to an identifiable production batch and its associated release documentation. The customer should be able to reconcile the material label, certificate of analysis, quantity, and relevant shipping records without ambiguity.
A certificate of analysis should contain, at minimum, an unambiguous product identification, batch number, applicable test results, specification limits, and an appropriate release authorization. Additional documents may be required depending on the customer's quality system, market, and contractual arrangements. These may include a safety data sheet, product specification, packaging details, or a quality questionnaire.
Document control is particularly important when specifications change. If an analytical method, test limit, manufacturing process, or packaging configuration is revised, the supplier should have a defined procedure for evaluating and communicating the change. Customers should establish which changes require prior notification or approval rather than relying on informal communication.
It is also important to distinguish between a supplier's company-level certifications and product-specific compliance. An ISO management system certification can provide evidence of a defined management framework, but it does not independently prove that a particular intermediate meets every customer specification or pharmaceutical regulatory requirement. Product release evidence and the scope of each certification should be reviewed separately.
Evaluating a Chinese Pharmaceutical Intermediate Supplier
China is an important sourcing market for pharmaceutical intermediates, but supplier selection should be based on demonstrated capability rather than geography alone. A qualified supplier should be able to explain its quality management arrangements, analytical resources, manufacturing controls, and approach to batch release.
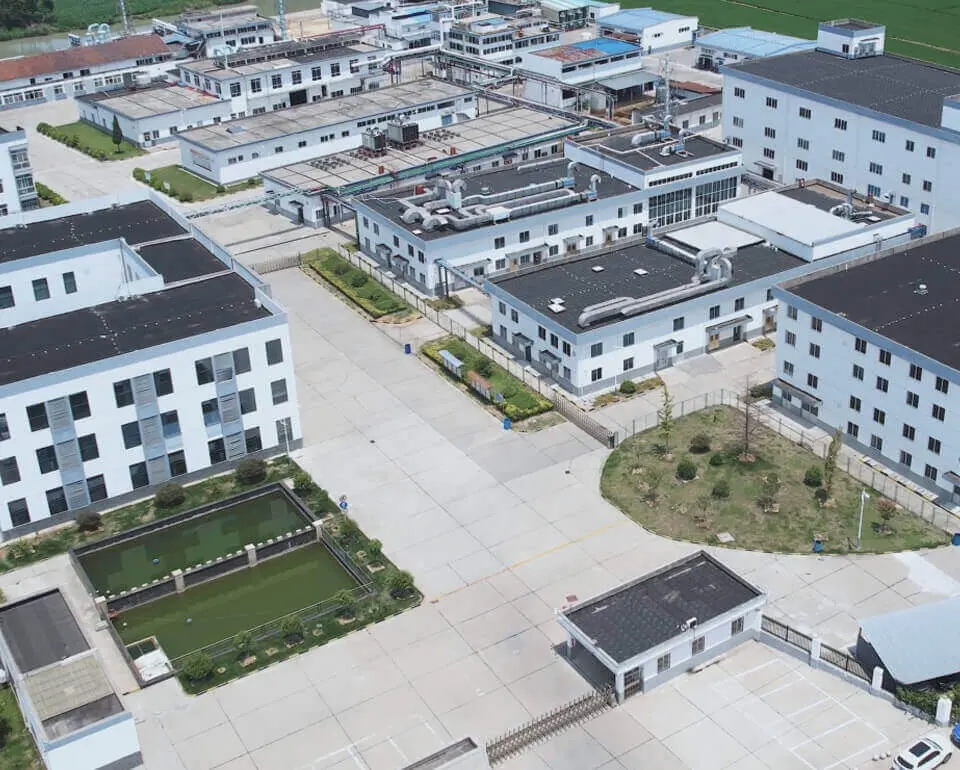
Jiangsu Jingye Pharmaceutical was founded in 1994 and restructured in 2016. The company operates an integrated business covering research and development, production, and international trade, with facilities covering 50,000 square meters in Changzhou, Jiangsu Province. Its technical capabilities include high-temperature and high-pressure reactions, hydrogenation, chiral synthesis, and high-vacuum distillation.
The company reports operating in accordance with GMP standards and maintaining modern production facilities, advanced analytical instruments, and an EHS management system. Its certifications include ISO 9001, ISO 14001, and GB/T 45001, alongside a Drug Production License. These company-level capabilities provide useful background when evaluating a potential manufacturing partner, while the actual product specification, manufacturing route, analytical methods, and release arrangements should still be confirmed for the specific intermediate.
Long-term cooperation with universities and research institutions can also support technical development and problem-solving. For a customer evaluating a new material source, the practical value lies in the supplier's ability to discuss technical requirements clearly, assess specification feasibility, and maintain reliable communication throughout qualification and supply.
Building a Practical Supplier Qualification Process
A supplier qualification process should begin with a technical review of the product specification and the documentation available for the proposed material. The customer can then assess whether the supplier's analytical methods address the required quality attributes and whether the reported results are sufficiently traceable.
Sample evaluation may provide additional evidence of suitability, but a laboratory sample should not automatically be treated as proof of commercial-scale consistency. Where the project progresses toward routine supply, the qualification process should consider manufacturing controls, change management, batch records, and the supplier's ability to maintain the agreed specification over time.
The review should also establish how deviations, out-of-specification results, complaints, and quality changes are investigated and communicated. These arrangements become especially important when material failures could interrupt downstream production or require costly investigations.
A clear division of responsibilities is beneficial. The supplier should provide accurate product information and appropriate release evidence, while the customer should define the material's intended use, required specification, and applicable qualification criteria. Where additional testing or documentation is needed, the requirements should be agreed before commercial delivery.
Conclusion
Choosing a 2-Bromo-1-[4-(methylsulfonyl)phenyl]-1-ethanone pharmaceutical intermediate supplier China requires a quality assessment that extends beyond the advertised purity figure. Structural confirmation, impurity profiling, method suitability, batch traceability, and change control all contribute to a more reliable evaluation of material quality.
Jiangsu Jingye Pharmaceutical offers an established pharmaceutical manufacturing background, technical capabilities across several specialized reaction and separation processes, and an international business footprint. For customers considering this specific intermediate, the next step is to confirm product-specific manufacturing and analytical capabilities, review the applicable specification, and establish clear quality documentation requirements before proceeding to routine supply.
A well-qualified supplier is not simply one that can provide a batch that meets today's test limits. It is one that can support a consistent, traceable, and technically defensible supply process as the customer's requirements evolve.